How Scientists Are Recharging T Cells Against Disease
In chronic viral infections and cancer, the killer cells often lapse into “exhausted” CD8+ T cells that can no longer stem the disease.
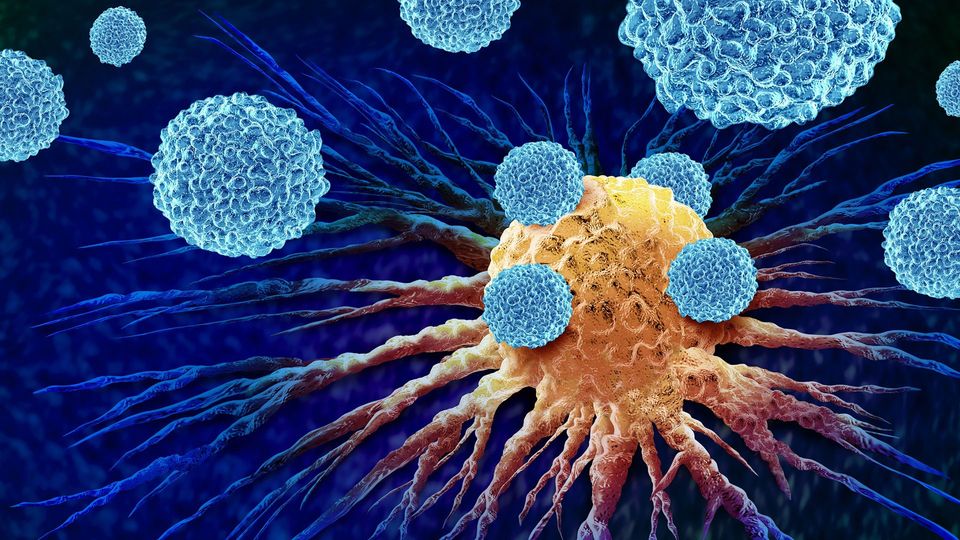
Killer immune cells destroy cancer cells and cells infected by virus. These CD8+ T cells are activated after detection of viral infection or growth of “non-self” tumor cells. However, in chronic viral infection and cancer, the killer cells often lapse into “exhausted” CD8+ T cells that no longer can stem disease.
This exhaustion is a major barrier in the new immunotherapies for cancer, including immune checkpoint blockers and CAR T cell therapy. In a detailed study of exhausted T cell subsets reported in Nature Communications, University of Alabama at Birmingham researchers led by Lewis Z. Shi, M.D., Ph.D., show that a transcriptional repressor called Gfi1, or growth factor independent-1, is a key regulator of the subset formation of exhausted CD8+ T cells and may offer a key to reducing exhaustion.
“Our study identifies an important role of Gfi1 in orchestrating CD8+ T cell response to anti-CTLA-4 therapy, the very first U.S. Food and Drug Administration-approved immune checkpoint blocker to treat patients with advanced cancer,” said Shi, a professor in the UAB Department of Radiation Oncology. “We reason that fine-tuning Gfi1 activity in T cells may prevent or reverse T cell exhaustion to bolster immune checkpoint blockade efficacy.”
Exhausted CD8+ T cells are a complex population of subsets composed of progenitor cells and “effector-like” or “terminally exhausted” cells. Effector-like cells still retain some killer ability. The UAB researchers used mice infected with a chronic virus to describe four subsets in the population, including a previously under-described Ly108+CX3CR1+ subset that expresses low levels of Gfi1, while other established subsets have high expression.
Notable key features of the Ly108+CX3CR1+ subset include: First, the Ly108+CX3CR1+ subset has a distinct chromatin profile from the other sets, meaning a changed accessibility to certain genes on the chromosome. Second, that subset is transitory and develops to terminally exhausted cells and effector-like cells, which retain some tumor killing ability. Third, this process depends on Gfi1.
To demonstrate a role for Gfi1 in immune checkpoint blockade therapy, the UAB team tested anti-CTLA-4 therapy in a mouse bladder cancer model, comparing mice that had T cells with either wild-type Gfi1 or Gfi1 knockout. They found that the anti-CTLA-4 therapy significantly inhibited tumor growth in wild-type but not Gfi1 knockout mice. Similarly, anti-CTLA-4 therapy promoted infiltration and/or expansion of CD4+ and CD8+ tumor-infiltrating lymphocytes in wild-type but not Gfi1 knockout mice. These observations were largely corroborated in a second mouse model of colorectal adenosarcoma, MC38.
“Considering Gfi1 downregulation is associated with the active differentiation of CD8+ T cell progenitors, we argue that transient and intermittent inhibition of Gfi1 with lysine-specific histone demethylase may facilitate the differentiation of progenitors to Ly108+CX3CR1+ cells and then to effector-like cells, thereby improving the control of chronic infections and tumors,” Shi said.
Along with a recent report by others of promising outcomes in small cell lung cancer from combining a lysine-specific histone demethylase inhibitor with the anti-PD-1 immune checkpoint blocker, “further testing of this combination approach should be conducted in melanoma, bladder cancer and colorectal adenocarcinoma, especially those resistant to immune checkpoint blockers,” Shi said.
Reference: Ojo OA, Shen H, Ingram JT, et al. Gfi1 controls the formation of effector-like CD8+ T cells during chronic infection and cancer. Nat Commun. 2025;16(1):4542. doi: 10.1038/s41467-025-59784-1
This article has been republished from the following materials. Note: material may have been edited for length and content. For further information, please contact the cited source. Our press release publishing policy can be accessed here.
 Add Technology Networks as a preferred Google source to see more of our trusted coverage.